





What Annex 1 requires on PUPSIT
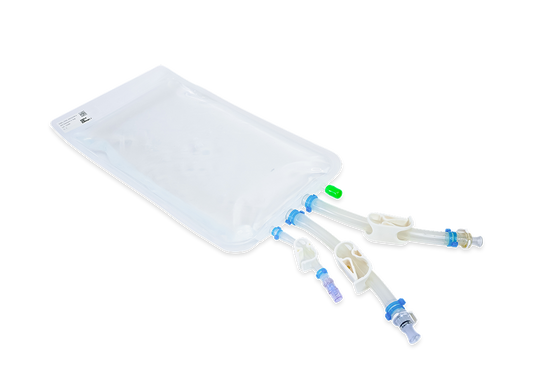
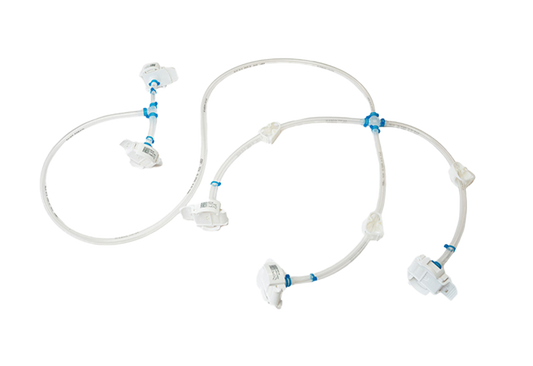
EU GMP Annex 1 basically treats pre‑use post‑sterilization integrity testing (PUPSIT) as a set of regulations to make sure sterilizing‑grade filters are still in perfect shape after they’ve been sterilized and before product use. If a manufacturer wants to skip it, they need a solid, risk‑based justification documented in their CCS/QRM strategy (contamination control strategy/ quality risk management). This article guides through the specific requirements of Annex 1 and sheds light on the diverse considerations with PUPSIT, including its methods, advantages, associated challenges, and the possibilities of single-use technologies. [[ToC]] What is Annex 1? EU GMP Annex 1 refers to a crucial document concerning pharmaceutical manufacturing. It serves as a comprehensive guideline, outlining the fundamental requirements and standards for the production of sterile medicinal products. Annex 1 is specifically focused on ensuring the quality, safety, and efficacy of sterile pharmaceuticals, which are critical for patient health and safety. This regulatory document provides detailed guidance on various aspects of sterile product manufacturing, including but not limited to facility design, personnel qualifications, process validation, equipment maintenance, and quality control measures. Annex 1 sets forth stringent criteria that manufacturers must adhere to in order to maintain sterility throughout the production process and minimize the risk of contamination or impurities in the final product. cGMP - everything you need to know Requirements of Annex 1 – what, where, by whom? Annex 1 delineates specific requirements that pharmaceutical manufacturers must adhere to in order to ensure the quality and safety of sterile medicinal products. These requirements cover various aspects of the manufacturing process, ranging from facility design to personnel training. What does Annex 1 require? Annex 1 outlines requirements for facility design, including cleanroom classifications, personnel, layout, and environmental controls to prevent contamination. It specifies criteria for equipment design, and maintenance to ensure suitability for sterile product manufacturing and minimize the risk of product contamination. Annex 1 mandates stringent process controls and validation procedures to verify the effectiveness and consistency of manufacturing processes. This requires thorough documentation and record-keeping to facilitate traceability and accountability throughout the manufacturing process. The document also emphasizes the importance of personnel training and hygiene practices to prevent microbial contamination and ensure product quality. Where do Annex 1 requirements apply? Annex 1 requirements apply to several stages of sterile medicinal product manufacturing, from raw material handling to final product packaging. These requirements are applicable to pharmaceutical manufacturing facilities, including cleanrooms, equipment, and utilities involved in sterile product production. Similarly to the FDA’s “Guidance for Industry: Sterile Drug Products Produced by Aseptic Processing Current Good Manufacturing Practice”, Annex 1 as issued by the European Union is a multinational standard. There are considerable similarities between these two frameworks; for instance, both determine PUPSIT as an essential step in ensuring process integrity in pharmaceutical manufacturing. By whom are Annex 1 requirements enforced? Annex 1 is enforced by the European Union, which has recently worked out a revision. A first draft of this revision was introduced in 2017, with the final version being published in 2022. By August 25, 2023, the revision of Annex 1 entered into force. Pharmaceutical manufacturers are responsible for ensuring compliance with Annex 1 guidelines within their facilities and operations. Quality assurance and regulatory affairs teams within pharmaceutical companies play a vital role in interpreting Annex 1 requirements, implementing necessary measures, and maintaining compliance with regulatory standards. EU (EMA / Annex 1) US (FDA) PUPSIT expected by default. Exceptions require a written, risk-based justification within CCS (contamination control strategy) Emphasis on post-use integrity testing for batch release. PUPSIT not universally mandated. "The integrity of the sterilized filter assembly should be verified by Integrity testing before use (PUPSIT)." Production and specific technologies - 8.87 "Integrity testing of the filter(s) can be performed prior to processing and should be routinely performed post-use." Sterile Drug Products Produced by Aseptic Processing - cGMP, Chapter IX PUPSIT – an overview Pre-use Post Sterilization Integrity Test (PUPSIT) is a crucial procedure in pharmaceutical manufacturing. It involves testing sterile filters before their use to ensure their integrity post-sterilization. The primary goal is to detect any potential defects or breaches that could compromise sterility during drug production. “The integrity of the sterilised filter assembly should be verified by integrity testing before use (pre-use post sterilisation integrity test or PUPSIT), to check for damage and loss of integrity caused by the filter preparation prior to use."EU GMP Annex 1 Paragraph 8.87 PUPSIT contributes to early identification of filter failures, enhances product quality, and aligns with regulatory standards. Mandated by agencies like the European Medicines Agency and the U.S. Food and Drug Administration, PUPSIT is integral in maintaining the reliability of sterile filtration systems, safeguarding pharmaceutical processes, and ensuring product safety and efficacy.[[1]] PUPSIT = Pre-Use Post-Sterilization Integrity Testing Integrity test performed after sterilization and prior to filtration for sterilizing-grade membrane filters used in aseptic processing. Performing PUPSIT – the 3 main methods Executing Pre-use Post Sterilization Integrity Test (PUPSIT) involves the application of three fundamental methods, each designed to assess the integrity of sterile filters before their integration into pharmaceutical manufacturing processes. 1. Bubble point testing – With the bubble point test, the pressure required to force a wetting liquid through the filter's pores is measured. This approach helps identify larger defects by observing the pressure at which the liquid begins to penetrate the filter. It is particularly effective in detecting significant breaches in filter integrity. 2. Diffusion testing – Utilizing the diffusion of a gas through the filter material, diffusion testing is a highly sensitive method for identifying smaller defects. This method is particularly effective in detecting minute breaches in filter integrity that might not be visible through other means. 3. Pressure hold testing – This method involves measuring the drop in pressure over time to identify leaks. Pressure hold testing is effective in detecting sustained leaks or breaches that might not be immediately apparent. It provides insights into the filter's ability to maintain pressure, contributing to a comprehensive assessment of filter integrity. Advantages of Pre-use Post Sterilization Integrity Tests By facilitating early detection of potential filter failures, PUPSIT offers a proactive approach to minimize the risk of compromised sterility during drug production. This contributes to the overall quality of pharmaceutical products, maintaining their purity and efficacy. Compliance with regulatory standards, mandated by entities such as the European Medicines Agency (EMA) and the U.S. Food and Drug Administration (FDA), is ensured through the implementation of PUPSIT as part of good manufacturing practices (GMP). PUPSIT serves as an essential component in validating sterile filtration systems, providing assurance that filters are free from defects and supporting the overall sterility of pharmaceutical manufacturing processes. By conducting PUPSIT, manufacturers effectively minimize the risk of potential contamination, ensuring sterile filters maintain an aseptic environment throughout the drug production process. Challenges when performing PUPSIT Performing Pre-use Post Sterilization Integrity Tests (PUPSIT) introduces several challenges, notably the intricate task of detecting minuscule defects in sterile filters. Despite the sensitivity of PUPSIT methods, identifying these tiny breaches can be challenging. Moreover, the necessity for filters to remain intact during testing adds another layer of complexity. The diverse array of filter types and materials further complicates testing approaches, requiring tailored strategies. Additionally, the time-sensitive nature of pharmaceutical production demands efficient PUPSIT execution without compromising thoroughness, posing an ongoing challenge. Striking a balance between rigorous testing and adhering to production timelines is crucial. Lastly, ensuring the seamless integration of PUPSIT within existing manufacturing processes without causing disruptions remains a key challenge for manufacturers. Filter integrity testing – single-use vs. multi-use In pharmaceutical manufacturing, the choice between single-use and multi-use filters brings several implications, also affecting PUPSIT. Multi-use filters, designed for repeated utilization, pose challenges for PUPSIT. The necessity for thorough cleaning between uses introduces complexities in filter validation, as procedures like autoclaving might affect their integrity. Conversely, single-use filters, tailored for one-time application, provide advantages in PUPSIT by ensuring a fresh, uncontaminated start for each test. Avoiding potential residue from previous use and in-house sterilization optimizes PUPSIT results, as sterilization is performed at the supplier. This dependency from the supplier, however, will be diminished with the emergence of x-ray sterilization as an increasingly popular approach. Automation and end-to-end fluid management systems to comply with Annex 1 The integration of automation and end-to-end fluid management systems stands as a crucial pillar in meeting the stringent standards set forth by GMP Annex 1 and its recent revision. Automation not only enhances operational efficiency but also plays a pivotal role in ensuring the sterility and integrity of the fluid management processes, as manual processes and thus the risk of human error can be reduced. This approach aligns seamlessly with Annex 1's emphasis on minimizing manual interventions, reducing contamination risks, and upholding the highest standards of product quality and patient safety. As the industry increasingly moves towards advanced end-to-end solutions, the synergy of automation and closed systems emerges as a cornerstone in achieving and sustaining compliance with GMP Annex 1. More and more pharmaceutical companies, CDMOs, and CMOs rely on specialized partners to establish future-proof manufacturing workflows that comply with current Good Manufacturing Practices. Annex 1 and PUPSIT requirements – partnering with Single Use Support Especially since the latest version of EU GMP Annex 1 has been issued, many insecurities have risen in the pharmaceutical sector – especially with regards to PUPSIT. Single Use Support is committed to ensure that the company’s fluid management solutions are compliant with Annex 1, as it allows the integration of PUPSIT directly into the closed fluid management environment, using their preferred filter integrity testing method. By realizing fully automated end-to-end processes with systems like the fluid management platform RoSS.FILL and its related single-use assemblies, pharmaceutical companies can significantly reduce the risk of human error. Furthermore, these scalable solutions address the urge of regulatory authorities to limit manual intervention in sterile fluid management. Product safety can be enhanced while simultaneously maximizing standardization, accuracy, transparency, and efficiency. Discover biopharma fluid transfer solutions References Pre-Use Post Sterilisation Integrity Testing (PUPSIT) in the new EU GMP Annex 1, https://www.gmp-compliance.org/gmp-news/pre-use-post-sterilisation-integrity-testing-pupsit-in-the-new-eu-gmp-annex-1, Published
READ MORE
